Two Etomidate API lots can both report a high assay result and still differ entirely in individual related substances, opposite enantiomer content, solvent residues, trace elements, or unknown peaks.
Etomidate is a chemically synthesized, single-enantiomer active pharmaceutical ingredient intended for parenteral drug products. This clinical application means stereochemical control, route dependent trace elements, and analytical method sensitivity are absolute requirements for source selection.
Evaluating a supplier on a simple specification sheet risks approving material with uncontrolled potent degradants, unstable profiles, or incompatible elements. Buyers need evidence that the manufacturer understands impurity sources, fate, detection, and control. This requires a thorough review of route informed risk assessments, named organic impurities, chiral controls, solvent and elemental data, nitrosamine evaluations, and multi batch stability trends.
Read this technical guide to establish a data driven impurity review before committing to an Etomidate source.
Why Assay And A Headline Purity Percentage Do Not Describe Impurity Risk
Assay, Chromatographic Purity, And Impurity Profile Answer Different Questions
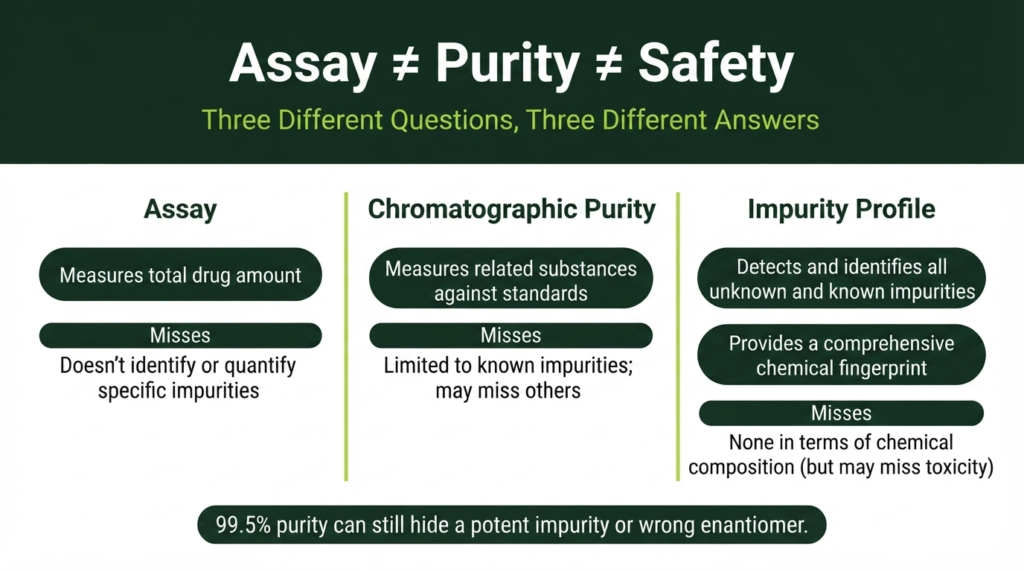
Assay measures the amount of Etomidate present relative to the stated basis. The result depends on the analytical method used and whether the value is calculated on a dried, anhydrous, or as is basis. Chromatographic purity is the relative detector response of peaks measured under one specific chromatographic procedure. This percentage is not equivalent to mass percent and ignores compounds the specific method cannot detect.
The impurity profile defines the identities, analytical descriptors, individual levels, total level, sources, and behavior of actual impurities. Equal peak areas do not mean equal mass when an impurity and Etomidate have different response factors. A 99.5 percent purity score can hide a highly potent impurity, an unresolved opposite enantiomer, or a rising degradant.
An Impurity Result Is Meaningful Only In Relation To Dose, Route, Method, And Intended Use
A percentage limit translates into an exposure risk only when paired with the maximum daily dose of the final drug product. The daily impurity exposure is calculated by multiplying the maximum daily API dose by the impurity fraction.
Parenteral use drives the exposure limits for Etomidate. Compendial conformance is the baseline requirement. An approved regulatory application or a buyer specific control strategy often requires additional named impurities, lower limits, or orthogonal testing.
| Evidence Label | What It Establishes | What It Does Not Establish | Buyer Follow Up |
| Assay | Amount of Etomidate by the stated method and basis | Identity and control of each impurity | Ask for impurity specific results and method scope |
| Related substances result | Peaks detected by that specific procedure | Chiral purity, solvents, elements, or nitrosamines | Map each risk to the appropriate analytical procedure |
| Total impurities | Sum defined by the method and reporting rules | Safety of each component or absence of potent impurities | Review named, unknown, and special risk impurities separately |
| “Complies” | Result met the cited acceptance criterion | Actual value, trend, or margin to limit | Obtain numerical data from recent batches |
Evaluating these quality documents requires understanding how to read an Etomidate certificate of analysis to separate pass and fail statements from trendable numerical data.
Build The Etomidate Impurity Map Before Reviewing Limits
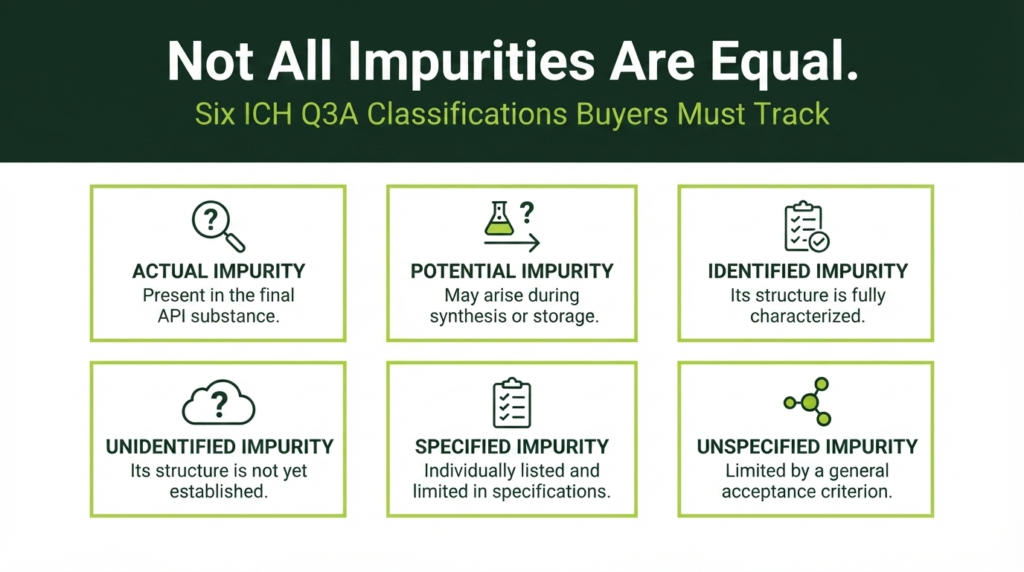
Separate Actual, Potential, Identified, Unidentified, Specified, And Unspecified Impurities
Classifying impurities using ICH Q3A definitions structures the technical evaluation. Buyers must confirm no plausible high risk impurity is ignored simply because it does not appear in the routine release table.
| Impurity Category | ICH Q3A Definition And Application | Buyer Decision Consequence |
| Actual | Appears in development, stability, validation, or commercial batches. | Trend and control it in routine testing. |
| Potential | Plausible from route chemistry, materials, reagents, or storage. | Demonstrate purge, absence, or ongoing control. |
| Identified | Chemical structure is characterized. | Evaluate source, response factor, fate, and safety. |
| Unidentified | Structure is not established. | Track by a reproducible descriptor such as relative retention time. |
| Specified | Individually named or designated with its own acceptance criterion. | Verify the specific limit against safety and capability data. |
| Unspecified | Controlled only by a general acceptance criterion. | Investigate when levels approach regulatory thresholds. |
Organize Risks By Source, Fate, And Control
The route informed assessment covers starting material carryover, reaction by products, reagent residues, stereochemical impurities, and degradation during processing or storage. For every potential impurity, request the source, formation point, purge step, analytical control, and supporting data. Theoretical purge arguments fall short when an impurity forms late in the process or shares chemical properties with Etomidate.
Etomidate Specific Organic Impurities Buyers Should Recognize
Desethyl-etomidate (Etomidate Acid) And Metomidate Are Relevant Named Reference Points
Etomidate is the ethyl ester of a chiral imidazole carboxylic acid. Desethyl-etomidate, also known as Etomidate acid, is the corresponding carboxylic acid formed after the loss or hydrolysis of the ethyl ester group. Metomidate is the corresponding methyl ester.
A database relationship or the availability of a reference standard establishes analytical relevance. Do not assume every commercial batch contains these specific compounds. Match the impurity standard by exact structure and stereochemistry rather than relying on the name alone. FDA GSRS records desethyl-etomidate entries with distinct stereochemical descriptions, requiring exact matching.
Treat Unknown Peaks As Information, Not Empty Space
Ask the supplier if unknown peaks are consistently tracked by relative retention time and whether the same unknown recurs across lots. Determine if a new peak appeared following a material, scale, or route change. Commercial process and stability impurities above identification thresholds require identification per ICH Q3A.
Names are not identities. Have an analytical reviewer verify the structure, salt form, stereochemistry, reference standard source, and intended analytical use before accepting any named impurity on a specification document.
| Etomidate Related Signal | Scientifically Supportable Context | Evidence To Request | Do Not Assume |
| Desethyl-etomidate / Etomidate acid | Structurally related carboxylic acid; plausible hydrolysis or process related impurity | Qualified standard identity, separation, response factor, release and stability trend | That it is always present or that storage is its only source |
| Metomidate | Structurally related methyl ester; USP uses a metomidate reference standard | Reference standard identity, method resolution, batch data, route rationale | That its source is identical for every manufacturer |
| Recurring unknown at stated RRT | Actual unidentified signal | Numeric trend, investigation status, method version, chromatograms | That “unknown” means uncontrolled or automatically unsafe |
| Route specific potential impurity | Plausible from a particular synthesis or material | Formation, fate, purge rationale, and confirmatory data | That public impurity catalogues are a complete route map |
Evaluate The Opposite Enantiomer Separately From Ordinary Related Substances
Why Etomidate’s Single Stereocenter Changes The Analytical Question
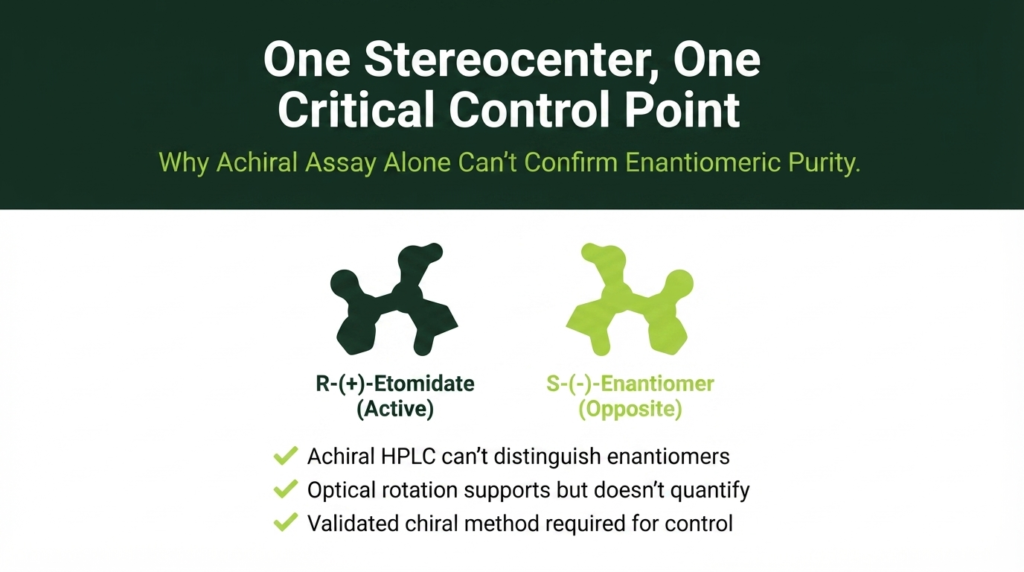
Marketed Etomidate is the R-(+)-enantiomer. Enantiomers have identical elemental compositions and behave identically in standard achiral HPLC methods. A high achiral assay cannot prove a low opposite enantiomer content. ICH Q3A explicitly excludes enantiomeric impurities from its scope. ICH Q6A governs chiral identity and opposite enantiomer control for a single enantiomer drug substance.
What Evidence Supports Stereochemical Control
Evaluate the chiral control strategy rather than demanding one specific test in isolation. Review where stereochemistry is introduced and whether later steps can cause racemization. An achiral assay combined with a separate opposite enantiomer control meets ICH Q6A allowances. Evaluate the chiral method for enantioselectivity, accuracy using the opposite enantiomer standard, and precision at low levels. Optical rotation results support overall stereochemical consistency but do not replace a validated chiral impurity procedure.
Review Residual Solvents From The Actual Process
Translate The Solvent Statement Into A Route-Informed Risk Assessment
ICH Q3C directs manufacturers to test for solvents used or reasonably likely to remain in the product. Classify the relevant solvents by Q3C class and determine whether control uses an Option 1 concentration limit or an Option 2 daily dose calculation. Include recovered solvents and solvent entrainment in wet cake or crystal lattices as possible risk inputs.
What A Defensible Residual Solvent Data Package Contains
Request a route-specific solvent list, specification limits, recent numerical results, and headspace GC method specificity. Identifying “Class 3 solvents only” is incomplete unless the specific solvents are named and confirmed. Understand how unspecified peaks in the GC chromatogram are handled. Environmental conditions impact solvent retention, a factor detailed in Etomidate API packaging and storage considerations.
Assess Elemental Impurities For Etomidate’s Parenteral Use
Start With Sources Of Elements, Then Apply The Correct Route Specific PDE
ICH Q3D establishes permitted daily exposures based on the final drug product. Parenteral applications require evaluation of potential Class 1, Class 2A, and relevant Class 3 elements. Map possible sources including intentionally added catalysts, water, manufacturing equipment, and container closure components. Do not apply oral PDEs to parenteral Etomidate assessments.
Separate A Risk Assessment Statement From Measured Evidence
A general declaration provides little value without route detail. Request representative multi batch ICP-MS or ICP-OES data. Convert any proposed API concentration limit to a daily exposure using the product maximum daily API dose. Confirm observed values are stable across representative lots rather than treating “below specification” as the end of the review.
Handle Mutagenic And Nitrosamine Risks As Separate, Low Level Questions
Apply ICH M7 To DNA Reactive Impurity Hazards, Not The Ordinary Q3A Threshold Logic
ICH M7 governs DNA reactive mutagenic impurities expected in the final drug substance. Acceptable levels are much lower than routine organic impurity thresholds. Identify route related structures and search bacterial mutagenicity data. When data are absent, apply two complementary (Q)SAR methodologies followed by expert review. Passing a routine related substances method controls an M7 impurity only if the limit of quantitation is demonstrably suitable for the threshold.
Read Multi Batch And Stability Data As A Trend
Assemble A Comparable Data Set
Request a table of at least three recent commercial or commercial representative batches. Include batch size, manufacture date, method version, individual named impurity values, recurring unknowns, total impurities, chiral impurity, relevant solvents, and stability time points. Demand numerical data for all trendable attributes.
Look For Direction, Discontinuities, And Method Artifacts
Evaluate the margin between the actual result and the control limit. Differentiate normal lot to lot spread from an unstable distribution. Investigate upward movement on stability or step changes after material shifts. A new impurity peak might simply reflect a lower LOQ or improved analytical separation. Monitor these metrics to evaluate long-term supplier reliability effectively.
| Observation | Plausible Explanations To Investigate | Decision Before Approval |
| Named impurity near limit in multiple lots | Process capability centered too close to limit; analytical bias | Request capability rationale and control improvement |
| Unknown peak grows on stability | Degradation, interaction, or method issue | Review identity threshold and stability indication |
| Abrupt profile shift | Route, raw material, site, scale, or method change | Establish comparability and change history |
| Total impurities stable but one component rises | Offsetting impurity changes hidden by total | Review individual profiles, not total alone |
Turn The Impurity Review Into An Approval Decision
Minimum Evidence To Request Before Technical Acceptance
Gather a consolidated set of documents to request from an Etomidate API supplier. This file must include specific datasets to enable a full quality risk assessment. Confidential route details often fall under the regulatory documents to evaluate when sourcing Etomidate API.
| Evidence Category | Required Documents And Data |
| Organic Impurities | Route informed risk assessment with source, fate, and purge rationale. |
| Named Standards | Identity and analytical role of named impurities, including stereochemistry. |
| Method Summaries | Current related substances and chiral control methods with validation performance. |
| Residual Solvents | Risk assessment tied to actual manufacture, with numerical data for relevant solvents. |
| Trace Elements | Risk assessment and representative target element data suitable for a parenteral product. |
| Mutagenic Risk | ICH M7 and FDA aligned nitrosamine risk assessments, with confirmatory results if warranted. |
| Batch Trends | Comparable recent batch and stability impurity trends, method versions, and excursion explanations. |
| Change Notification | Commitment for impurity relevant route, material, site, method, or packaging changes. |
Use Three Outcomes Instead Of A Simplistic Pass Or Fail
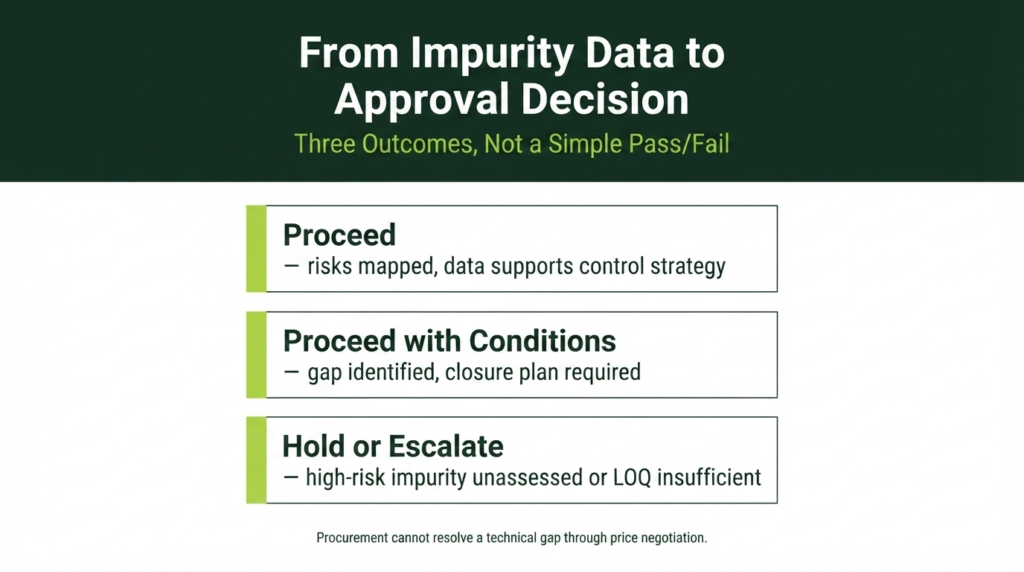
| Approval Outcome | Definition | Next Steps |
| Proceed | Risks are mapped, methods quantify at relevant levels, and data fits the product control strategy. | Approve the source for commercial supply. |
| Proceed with targeted conditions | Evidence is adequate but a defined gap has a dedicated closure plan. | Resolve the gap before use in commercial manufacturing. |
| Hold or escalate | High risk impurities lack assessment or method LOQs exceed required limits. | Reject the material or escalate to quality leadership. |
Procurement teams must not solve a technical gap through price negotiation. The quality owner determines whether the evidence supports safe use.
Share your intended market, dosage form, maximum daily dose, regulatory basis, and required impurity controls with Velcare to request an accurate Etomidate API quote.
Questions Buyers Frequently Ask
Is 99 Percent Or 99.5 Percent Etomidate API Purity Enough For Pharmaceutical Use?
No. The distribution and identity of the remainder, chiral purity, method coverage, and patient exposure determine suitability. A high purity percentage is meaningless if it obscures a mutagenic impurity or the wrong enantiomer.
Are All Etomidate Impurities Listed On The COA?
No. A certificate of analysis reports tests defined by the current specification and reporting conventions. Potential impurities, sub reporting threshold signals, and route risk assessments sit in the broader quality package.
Is Metomidate The Same As Etomidate Acid?
No. Metomidate is the methyl ester analogue. Desethyl-etomidate, known as Etomidate acid, is the corresponding carboxylic acid. Require exact structures and stereochemical forms when ordering standards or interpreting peaks.
Does USP Compliance Eliminate The Need For Nitrosamine And Elemental Assessments?
No. Compendial testing does not replace current route and product specific ICH risk assessments or the control strategy enforced by the drug product sponsor.
Should A Buyer Demand Zero Impurities?
No. Zero is analytically undefined without a known detection capability. Controls reflect safety data, process capability, and fit for purpose methods. A non detect result means the compound is below a stated quantitation level.